My contribution
I was responsible for the computational part of an ongoing joint study of concentrated DME electrolytes. I assessed classical force-field limitations, developed and validated a fine-tuned MACE model, and generated and analyzed molecular-dynamics trajectories.
I used PCA of model descriptors to select diverse training structures, generated DFT reference data, and used the fine-tuned model to study coordination, ionic-network dynamics, and transport. This is fine-tuning and application of MACE, not authorship of the original MACE foundation model.
Validation
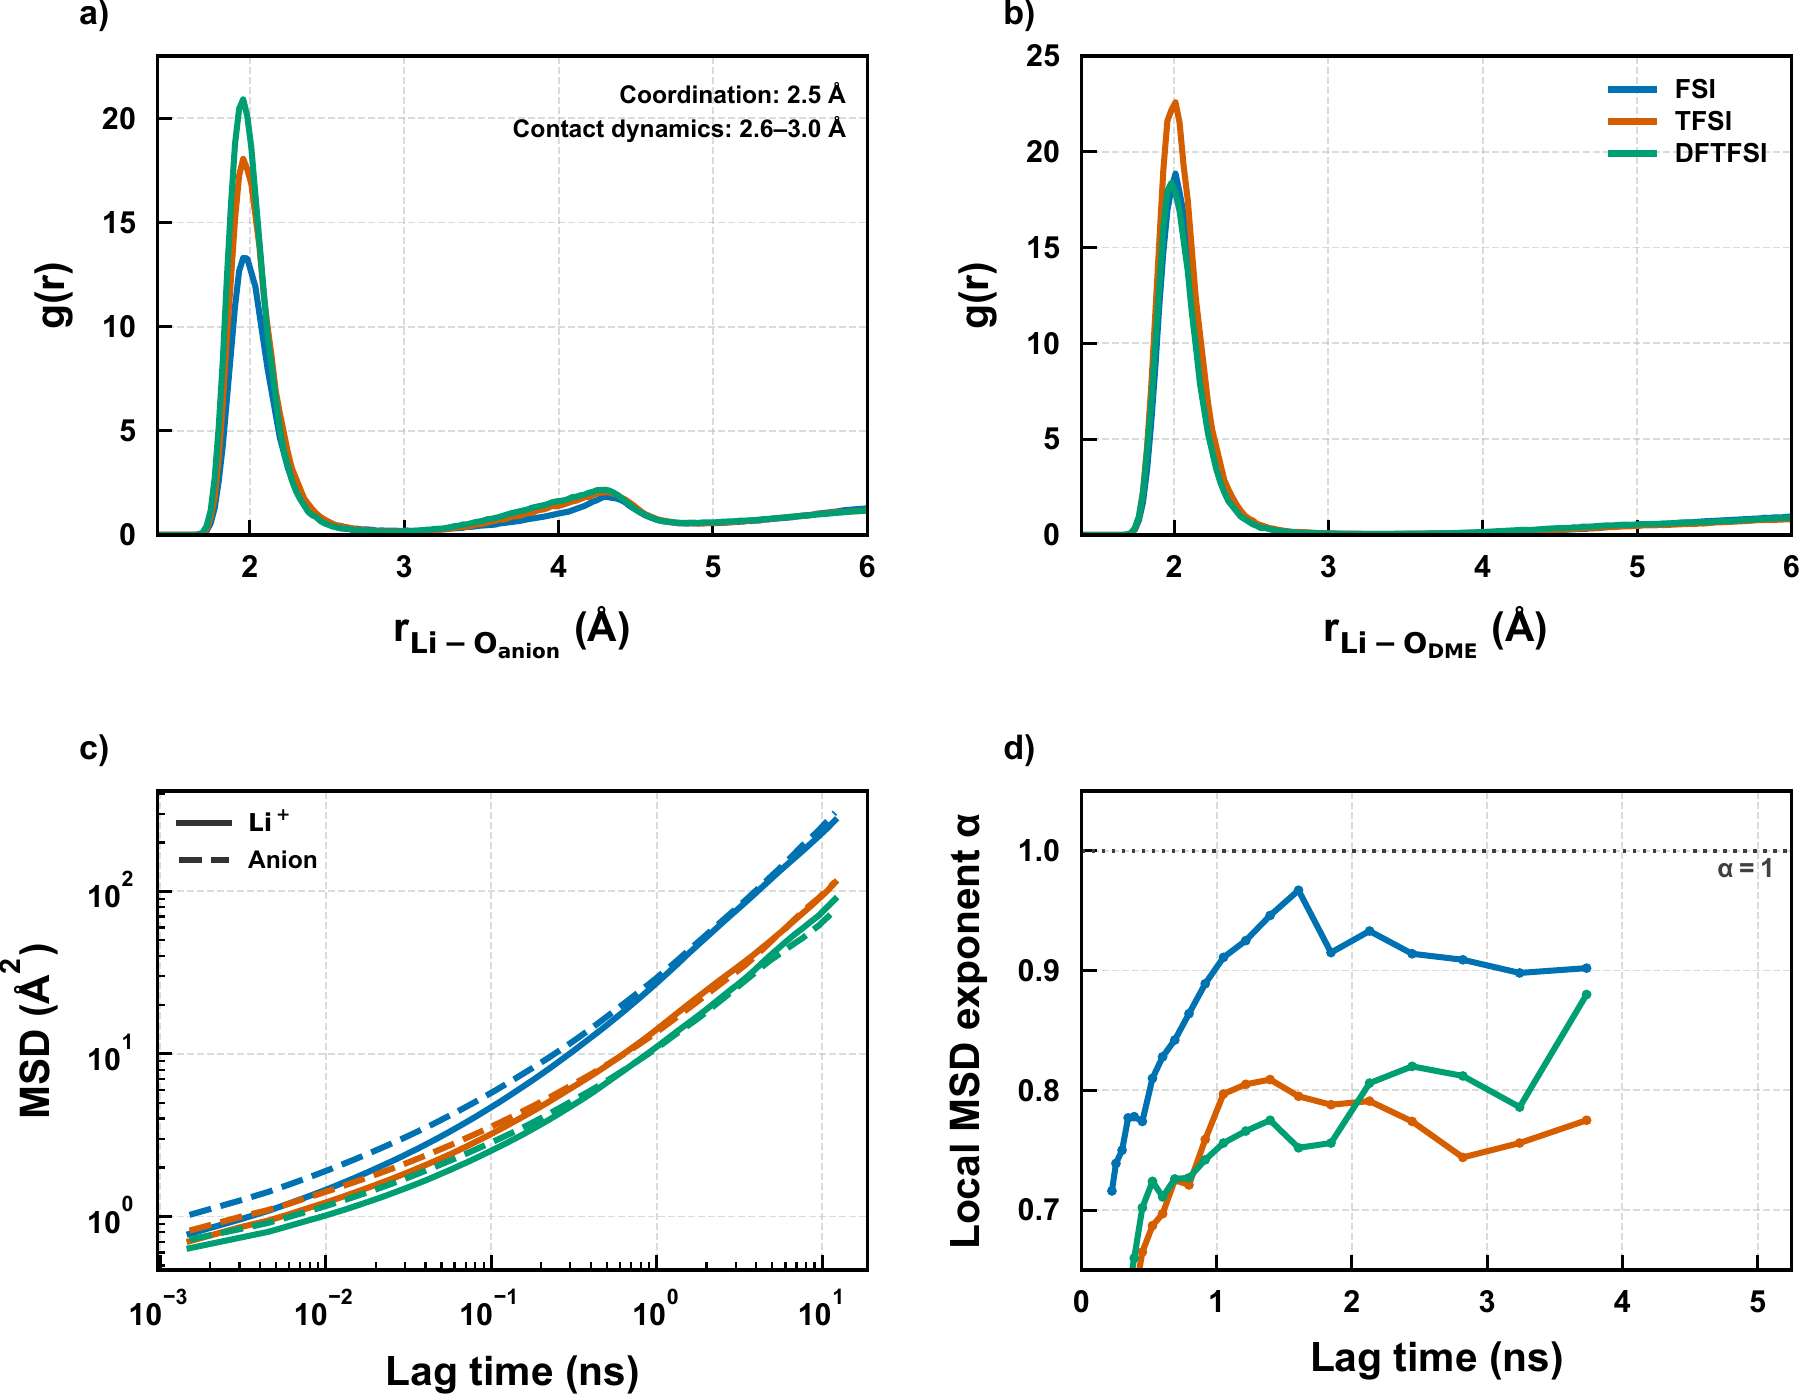
The diagnostic figure shows lithium–anion and lithium–DME radial distributions, mean-square displacements, and local MSD exponents at 300 K. These checks connect the learned potential to the behavior of the simulated liquid.
Some trajectories retain subdiffusive exponents at late lag times. I therefore treat the transport values as effective estimates over the sampled window, rather than fully converged long-time diffusion coefficients.
Research status
The joint manuscript on anion-dependent structure, transport, and interphase formation is in preparation. My role covers simulation, analysis, mechanistic interpretation, and manuscript development; the experimental program is led by collaborators.